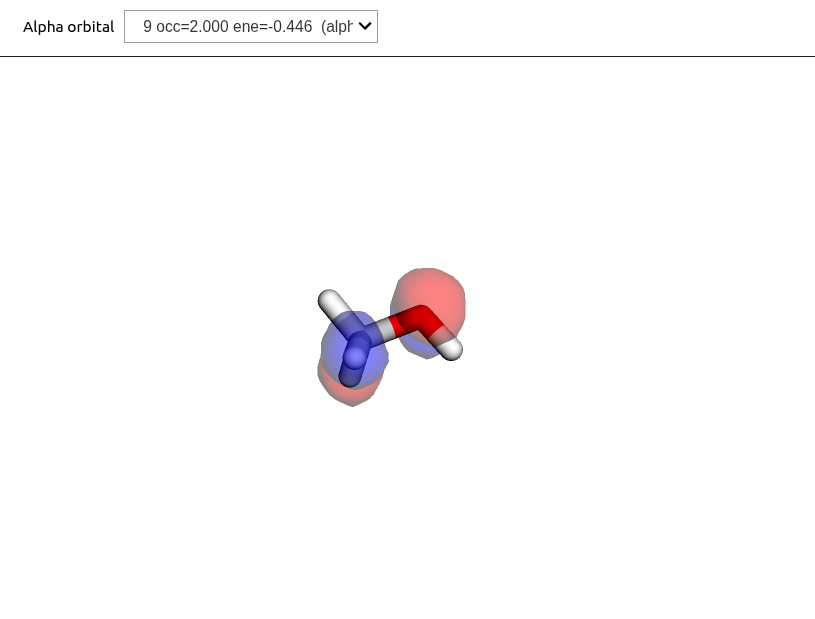
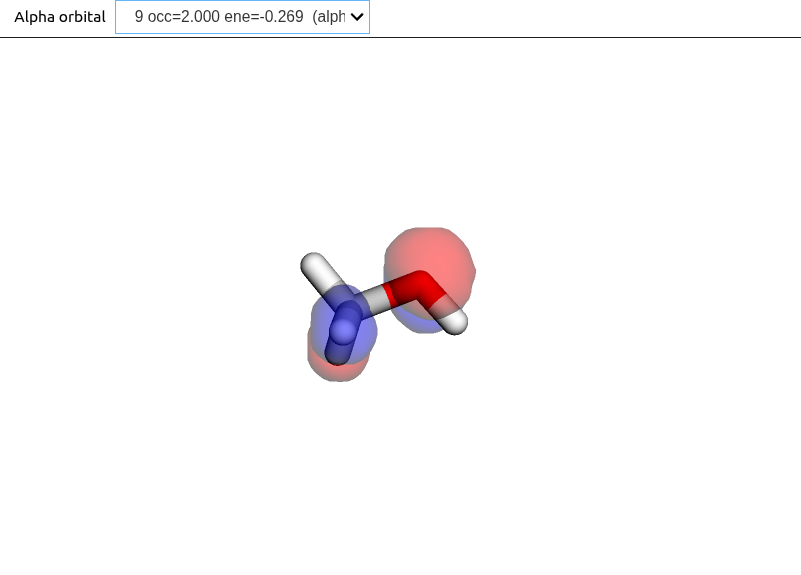
Here we consider the optimization of the electronic structure using Hartree--Fock (HF) and Kohn-Sham density functional theory (KS-DFT), and how to plot the resulting molecular orbitals. Our example is methanol, using the xTB structure from the previous section.
At the HF/6-311G level of theory, the electronic structure is optimized by creating the molecule and basis objects, setting up an unrestricted SCF driver, and performing the SCF calculation:
                                            Self Consistent Field Driver Setup
                                           ====================================

                   Wave Function Model             : Spin-Restricted Kohn-Sham
                   Initial Guess Model             : Superposition of Atomic Densities
                   Convergence Accelerator         : Two Level Direct Inversion of Iterative Subspace
                   Max. Number of Iterations       : 50
                   Max. Number of Error Vectors    : 10
                   Convergence Threshold           : 1.0e-06
                   ERI Screening Threshold         : 1.0e-12
                   Linear Dependence Threshold     : 1.0e-06
                   Exchange-Correlation Functional : B3LYP
                   Molecular Grid Level            : 4

* Info * Using the B3LYP functional.

         P. J. Stephens, F. J. Devlin, C. F. Chabalowski, and M. J. Frisch.,  J. Phys. Chem. 98, 11623 (1994)

* Info * Using the Libxc library (v7.0.0).

         S. Lehtola, C. Steigemann, M. J.T. Oliveira, and M. A.L. Marques.,  SoftwareX 7, 1–5 (2018)

* Info * Using the following algorithm for XC numerical integration.

         J. Kussmann, H. Laqua and C. Ochsenfeld, J. Chem. Theory Comput. 2021, 17, 1512-1521

* Info * Starting Reduced Basis SCF calculation...
* Info * ...done. SCF energy in reduced basis set: -115.018935737583 a.u. Time: 0.07 sec.


               Iter. |    Kohn-Sham Energy | Energy Change | Gradient Norm | Max. Gradient | Density Change
               --------------------------------------------------------------------------------------------
                  1      -115.712788803550    0.0000000000      0.12055772      0.01852446      0.00000000
                  2      -115.712969331741   -0.0001805282      0.10625363      0.01806455      0.05103107
                  3      -115.714365718827   -0.0013963871      0.00476261      0.00081966      0.02632569
                  4      -115.714368563845   -0.0000028450      0.00049882      0.00007594      0.00147822
                  5      -115.714368569914   -0.0000000061      0.00046205      0.00007408      0.00021746
                  6      -115.714368587631   -0.0000000177      0.00003830      0.00000547      0.00009628
                  7      -115.714368587831   -0.0000000002      0.00000282      0.00000036      0.00001735
                  8      -115.714368587833   -0.0000000000      0.00000034      0.00000006      0.00000156

               *** SCF converged in 8 iterations. Time: 0.20 sec.

               Spin-Restricted Kohn-Sham:
               --------------------------
               Total Energy                       :     -115.7143685878 a.u.
               Electronic Energy                  :     -156.2327230764 a.u.
               Nuclear Repulsion Energy           :       40.5183544886 a.u.
               ------------------------------------
               Gradient Norm                      :        0.0000003430 a.u.


               Ground State Information
               ------------------------
               Charge of Molecule            :  0.0
               Multiplicity (2S+1)           :  1
               Magnetic Quantum Number (M_S) :  0.0
print(f'The total (B3LYP) energy is {scf_b3lyp.get_scf_energy(): .5f} Hartree')